
IMPD Clinical Trials: key quality considerations for regulatory success
IMPD Clinical Trials: key quality considerations for regulatory success
Regulatory Affairs Pharma & Biotech
Wednesday, 30 April 2025

Stella Grigoriou, Specialist RA Pharma at QbD Group

 Share
 \
\
X (Twitter)
 \
\
LinkedIn
 \
\
WhatsApp
 \
\
Copy link
 \
\
Mail
Prepare a compliant IMPD for clinical trials. Avoid delays with expert tips on quality documentation, common pitfalls, and regulatory expectations.
Play
IMPD Clinical Trials: key quality considerations for regulatory success
AI-generated audio
9:17
An Investigational Medicinal Product Dossier (IMPD) is a critical document required for clinical trial applications (CTAs) in the European Union and the United Kingdom. It is governed by Regulation (EU) No 536/2014 in the EU and by The Medicines for Human Use (Clinical Trials) Regulations 2004 SI 2004 No. 1031 in the UK. The IMPD is generally organized into 4 primary sections:
- Quality
- Non-clinical
- Clinical
- Safety and efficacy
In this blog post, we focus on the key considerations for the Quality section of the IMPD for chemically defined investigational medicinal products (IMP).
After months of work on your CTA, ensuring your IMPD meets quality standards sets you up for success and avoids unnecessary delays. Common issues like insufficient justification of specifications or missing batch analysis data can lead to regulatory Requests for Information (RFIs), adding time and complexity to the review process.
Having worked on over 100 CTAs across the EU and UK, we’ve supported numerous companies in navigating these challenges and maintaining regulatory compliance. Preparing a thorough, compliant IMPD from the start helps avoid these setbacks, ensuring smoother progression to clinical phases and keeping your trial on track.
What is the IMPD quality section?
The IMPD quality section must provide comprehensive documentation on the chemical and pharmaceutical quality of the IMPs. This covers aspects like Chemistry, Manufacturing, and Controls (CMC), stability data and packaging.
The Quality IMPD structure aligns with the Common Technical Document (CTD) format, particularly CTD Module 3: Quality. The module headings in Figure 1 are used when writing the Quality IMPD for Drug Substance and Drug Product.
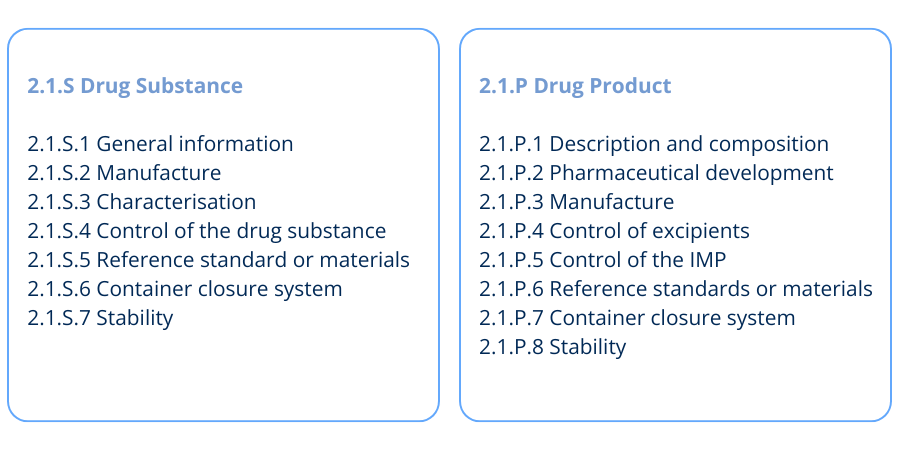 Figure 1 - Quality Module Headings for Drug Substance and Drug Product in IMPD
It’s also important to consider how requirements evolve throughout clinical development. In Phase I, the focus is on safety, with limited manufacturing and characterisation data.
By Phase III, however, regulators expect detailed information on stability, specifications, and comprehensive batch data (see Figure 2). This makes phase-specific planning essential for sponsors working with IMPDs in clinical trials.
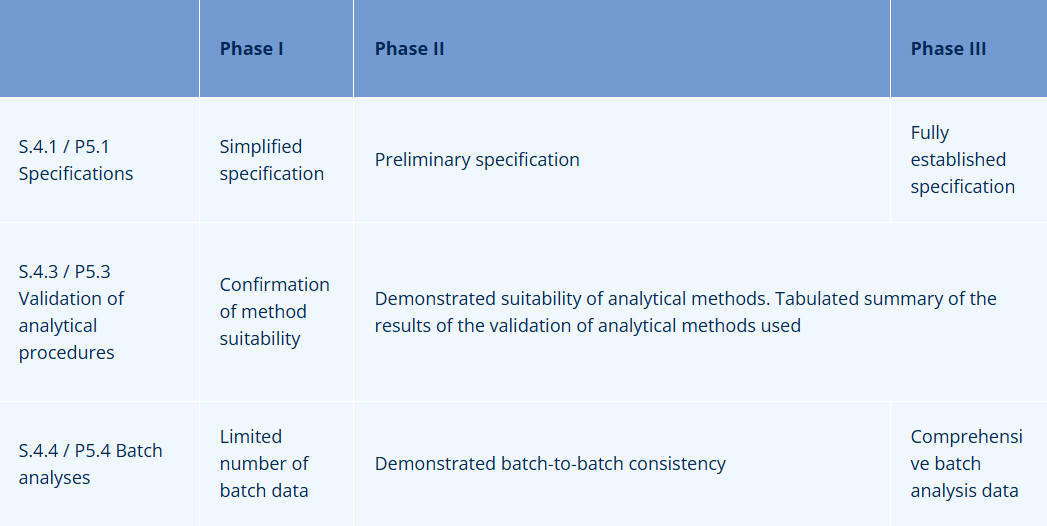
Table 1 - Increasing Regulatory Requirements for IMPD Quality Section from Phase I to Phase III
Key quality considerations for IMPD Clinical Trials
Below is a summary of key considerations for the requirements of each IMPD section. These are based on the common deficiencies identified by the Medicines and Healthcare products Regulatory Agency (MHRA) when assessing CTAs as well as QbD’s extensive experience in this area.
1. Characterisation data
Missing, inadequate, or poorly interpreted characterisation data is a frequent issue. Regulators typically expect the drug substance structure to be confirmed by at least two spectroscopic methods, with representative spectra included in the IMPD.
2. Method validation data
The IMPD must include a summary of analytical method validation for Phase II and III trials. In Phase I, confirmation of method suitability is sufficient. Inadequate summaries can result in deficiencies.
3. Batch analysis data
These data demonstrate that each manufacturer can consistently produce the drug substance and product at the proposed scale(s) throughout clinical development. Data must be included for all listed manufacturing sites and inclusion of drug substance batches used for non-clinical studies should not be overlooked. Missing this information can trigger RFIs.
4. Justification of specifications
Sponsors must justify specifications, acceptance criteria, and control methods. Impurity limits should be backed by safety/toxicity data and batch data. If expected characteristics are not controlled, this must be justified.
5. Retest period
Stability data should be summarised in table form, showing that the drug substance meets specifications at manufacture. The re-test period should be based on this data. Missing or insufficient stability data often results in RFIs.
6. Shelf life
The proposed shelf life (IMP) must be supported by stability data, although extrapolation can be usedto set this. A shelf-life extension plan and justification criteria must also be included. This is a common deficiency flagged by agencies.
7. TSE/BSE certification
Sponsors must include documentation controlling for transmissible spongiform encephalopathy (TSE) agents, such as certification and relevant process controls. Missing or incomplete data will result in RFIs.
8. Qualified Person (QP) declarations
Sites manufacturing IMP that are located outside of the EU/EEA require a QP declaration confirming EU GMP compliance. This will require an on-site GMP audit, so planning ahead is crucial. Incomplete declarations are a frequent source of delays.
9. Labelling
CTA submissions must include labelling samples for all IMPs, including comparators, in line with Annex 13 of EU GMP guidelines. Any deviations must be justified. Unjustified deviations often require clarification or amendment.
10. GMP Compliance
The IMPD must be supported by GMP documentation such as the Manufacturing and ImportationAuthorisation (MIA) and QP certification (Annex I, Section G, Clinical Trial Regulation (CTR). MIA is required for all manufacturing, packaging, and labelling sites.
Common IMPD mistakes to avoid in Clinical Trials
Transitioning between regulatory frameworks, such as from the IMPD to an IND (Investigational New Drug) application, can be challenging due to differences in content, format, and requirements. Comparing the two and maintaining alignment is critical.
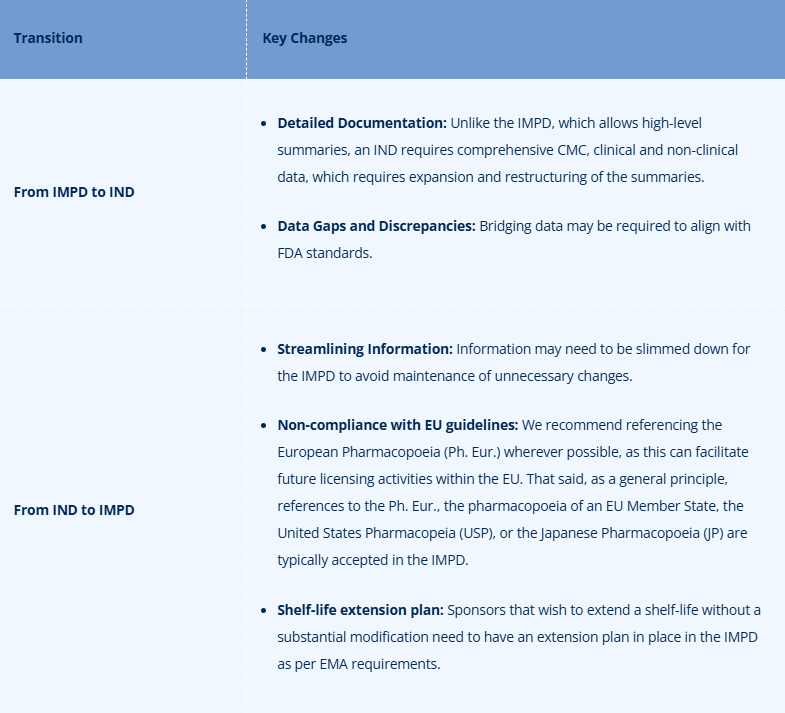
Table 2 - Key regulatory differences when transitioning between IMPD and IND submissions
Writing the IMPD quality section efficiently
A well-written IMPD Quality section can save time, prevent RFIs, and support smoother clinical trial progress. The following strategies help ensure your documentation is both compliant and efficient:
- Adhering to guidelines: Follow the European Commission's Guideline on the Requirements for the Chemical and Pharmaceutical Quality Documentation.
- Automation and templates: Streamline your IMPD preparation process with internal templates.
- Versioning: Maintain strong version control to reflect ongoing changes. This is particularly important for IMPD clinical trials involving multiple studies or regions.
Why work with experts for IMPD Clinical Trials?
It’s highly recommended to involve an IMPD subject matter expert before submission. They can help proactively identify and resolve potential gaps, ensuring compliance and minimizing delays.
QbD has supported numerous companies in obtaining and maintaining compliant CTAs in the EU and UK. Our expert team can assist with authoring, reviewing, and maintaining IMPDs in clinical trials, as well as responding to agency queries. With up-to-date regulatory intelligence and a database of common deficiencies, we ensure a smooth submission process.
Need support with your IMPD?
Reach out to QbD’s regulatory affairs experts today. We’ll help you prepare a strong, compliant IMPD so you can focus on moving your clinical trial forward with confidence.
Let’s talk about how we can support your team.
Regulatory Affairs Pharma and Biotech
We support you from concept to launch in the full lifecycle
References
1.EUR-Lex - 02014R0536-20221205 - EN - EUR-Lex [Internet]. eur-lex.europa.eu.
The Medicines for Human Use (Clinical Trials) Regulations 2004 [Internet]. Legislation.gov.uk. 2015.
UK Government. Common issues identified during clinical trial applications.
European Commission. Guideline on the requirements for the chemical and pharmaceutical quality documentation concerning investigational medicinal products in clinical trials.
Stay ahead in life sciences
Keeping up with the fast-paced life sciences industry doesn’t have to be overwhelming.
Our newsletter delivers the latest insights, industry updates, and expert content directly to your inbox, helping you stay informed and make smarter decisions.

Discover more expert content
Webinar
From Strategy to First-in-Human: A Practical Roadmap for Biotech Development
Join our webinar to learn how to navigate nitrosamine risks and streamline your biotech development from early planning to first-in-human trials. Register now for expert insights.
Webinar
PMS & PMCF Compliance for Medical Devices Under EU MDR
Gain practical insights on PMS & PMCF compliance for medical devices under EU MDR. Watch our webinars on demand.
Case study
A Decade of Excellence: Linguistic Review Support for a Global Pharma Leader
QbD Group streamlines linguistic reviews for a leading pharma client, ensuring compliance and accuracy across 25 EU/EEA languages while meeting stringent regulatory deadlines.
Case study
Supporting Market Access to Essential Medicines in Europe
Support your entry into the European pharmaceutical market with expert regulatory guidance, ensuring compliance and access to essential medicines for patients. Contact us today for assistance.

Whitepaper
Medical Device Regulation (MDR) Checklist
Implement the Medical Device Regulation (MDR) with ease. Download our checklist of mandatory documents for MDR compliance.
Whitepaper
Regulatory Affairs for Pharma and Biotech
In this flyer, you will learn more about the regulatory services QbD Group provides for the pharmaceutical and biotechnology industries.
Whitepaper
Clinical Evaluation for Medical Devices under MDR
This whitepaper will guide you through crucial regulatory documents pertaining to the clinical evaluation process of your medical device. Download now!
Whitepaper
IVDR extension: what does this mean for you?
In this whitepaper, you will learn the impact of the IVDR transition extension and receive tips and strategies to navigate these regulatory changes.
Webinar
Drug-Device Combination Products and Article 117
Explore the regulatory complexities of Drug-Device Combination Products. Learn about Article 117 requirements, Notified Body Opinions, and more.
Whitepaper
Mobile health on the rise: exploring the regulatory landscape for reimbursement
This whitepaper will help you navigate the maze of the DTx regulatory environment, highlighting several important countries and regulations.
Whitepaper
Clinical investigations under MDR: key insights and practical experiences
This whitepaper covers clinical investigation authorization under MDR, the EU Legal Representative's role, and the MDR's impact on study conduct and post-market surveillance.
Case study
CE Mark Renewal for bioXtra Products: A Success Story
Discover how Lifestream Pharma successfully renewed the CE mark for their BioXtra Dry Mouth products under the EU MDR. Learn about the challenges faced, the approach taken for compliance, and the results achieved.
Webinar
Technical Documentation Essentials for Medical Device Software
Gain practical insights on crafting MDR-compliant technical documentation for Medical Device Software.

Webinar
Getting Started: Overcoming Initial Obstacles in Medical Device Software Development
Watch our webinar on demand and learn about regulatory obstacles, MDR, AI Act, and best practices for medical device software development and market entry.
Webinar
Mastering Clinical Performance Studies under IVDR
In the evolving regulatory landscape of IVDs, demonstrating clinical performance is more crucial and challenging than ever. Discover more in this webinar on demand.
Whitepaper
Clinical evidence for In Vitro Diagnostics under IVDR
In this whitepaper, we will guide you through the documents required for the demonstration of clinical evidence under the IVDR. Download now.
Webinar
Post-Market Surveillance Requirements for Medical Devices and IVDs
Learn about post-market surveillance requirements under MDR and IVDR regulations. Gain insights from industry experts in this on-demand webinar.
Whitepaper
The key role of Regulatory Affairs in the pharmaceutical industry: from drug development to commercialization
Discover the key role of Regulatory Affairs in pharma and how Regulatory Affairs teams support product lifecycle management in this whitepaper. Download now.

Webinar
3D Printing in the Healthcare World
Explore the challenges and opportunities of 3D printing in healthcare, featuring expert insights on regulations, pitfalls, and efficiency.
Webinar
IVDR Extension Explained: Tips & Strategies
Gain valuable insights on the IVDR extension and its impact on IVD manufacturers with our webinar on demand.
Whitepaper
Innovations in ATMP: state of the industry in 2024
This whitepaper explores the innovations in ATMP and solutions in detail, with a focus on the current landscape as of April 2024. Download now.
Case study
The journey toward IVDR compliance for the LVOne device
QbD Group helped Upfront Diagnostics gain entry into the MD/IVD industry by supporting them on their path to ISO 13485 certification.
Case study
Ensuring a smooth MDR transition for Oystershell's medical devices
The landscape of medical device regulations is constantly evolving, making it a challenge for companies to keep up. Our collaboration with Oystershell, which began in July 2023, highlights our expertise in transforming complex challenges into success stories, all with a touch of QbD Clinical magic.
Case study
Successful clinical evaluation supports rapid CE marking of Minze Health's Uroflowmeter
QbD Clinical assisted Minze Health, a urology-focused medical devices company, in obtaining CE marking under MDR through clinical evaluation.

Webinar
The Medical Device CE Certification Pathway Explained: From Classification to Conformity Assessment by the Notified Body
Learn about the Medical Device CE Certification Pathway, from classification to conformity assessment by a Notified Body. Join our webinar on demand.

Case study
Helping Relu to comply with changing regulations
QbD Group supported software company Relu with their expertise to comply with changing regulations.

Case study
Quickly bringing a new COVID-19 medical device to market
Obtaining the required government certifications for a medical device is always a challenge. Especially when the COVID clock is ticking.
Twitter Widget Iframe