
Analytical Method Validation: are your analytical methods suitable for intended use?
Analytical Method Validation: are your analytical methods suitable for intended use?
Qualification & Validation Pharma & Biotech
Thursday, 13 April 2023

Claudia Wever, Project Manager at QbD Group

 Share
 \
\
X (Twitter)
 \
\
LinkedIn
 \
\
WhatsApp
 \
\
Copy link
 \
\
Mail
In this blog post, you will learn more about analytical method validation and the criteria to consider when validating your analytical method.
Play
Analytical Method Validation: are your analytical methods suitable for intended use? | QbD Group
AI-generated audio
6:59
What is Analytical Method Validation?
Analytical method validation is a critical process in the pharmaceutical, biotechnology, and food industries to ensure the quality and safety of products.
The objective of the validation of an analytical method is to demonstrate that it is suitable for its intended purpose. Both the method and its intended use should be clearly defined. This means the method must be fully developed and optimized before initiating method validation.
The following parameters must be known:
- What analytes are to be analyzed?
- What are their expected concentration levels?
- What is the sample matrix and is any interference expected?
- Should the method yield a quantitative or qualitative result?
In this blog post, we will give an overview of the criteria to consider when validating your analytical method.
Free expert article
Analytical Method Validation
In this publication, we will give an overview of the criteria to consider when validating your analytical method.
What makes an analytical method ‘suitable’ for intended use?
Now, what makes an analytical method ‘ suitable’? An analytical method is considered suitable when it has been established that the results generated by the method are consistent, reproducible, and reliable.
The parameters required (at minimum) to establish suitability are described in the ICH Q2a guideline: Validation of Analytical Procedures: Text and Methodology (See Table 1).
Note: it is not required that these tests be performed separately, so it is allowed to create a larger dataset to allow multiple parameters to be determined in one run.
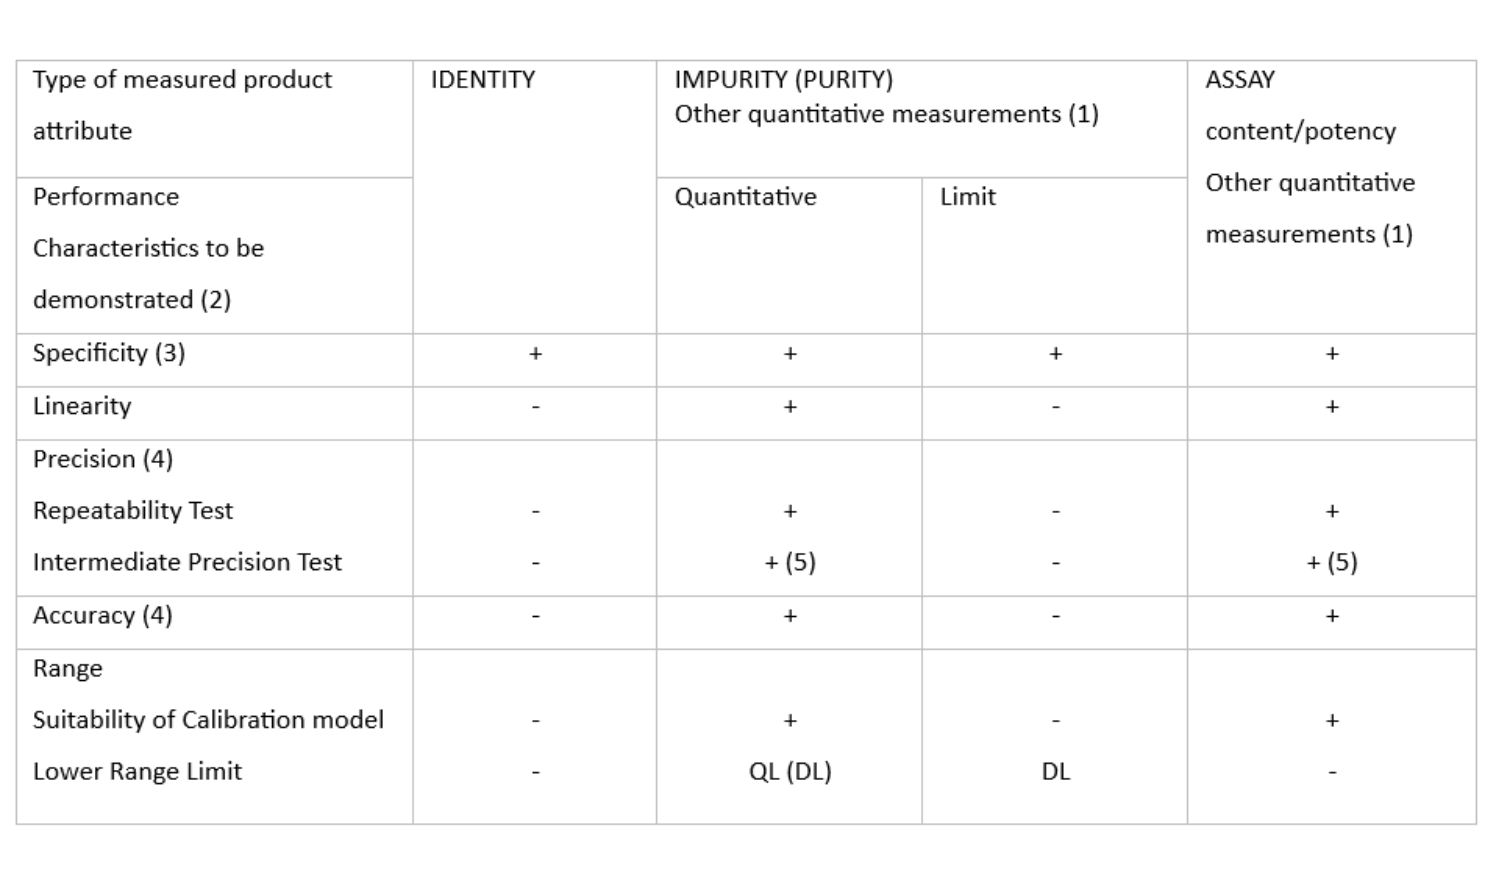
Table 1: Typical performance characteristics and related validation tests for measured product attributes (ICH Q2)
“-“ means that this test is not usually performed
“+” means that this test is usually performed
( ) means that this test is usually not performed, but in some complex cases recommended
QL, DL: Quantification Limit, Detection Limit
(1) other quantitative measurements can follow the scheme of impurity testing, if the working range is close to the detection or quantitation limits of the technology, otherwise following the assay scheme is recommended.
(2) some performance characteristics can be substituted with technology inherent justification or qualification in the case of certain analytical procedures for physicochemical properties.
(3) a combined approach can be used alternatively to evaluate accuracy and precision separately
(4) lack of specificity of one analytical procedure could be compensated by one or more other supporting analytical procedures
(5) Reproducibility and intermediate precision can be performed as a single set of experiments.
Typical analytical method validation characteristics
Specificity
Specificity is the ability of the method to measure only the analyte of interest without interference from other components in the sample that are likely to be present, such as impurities, degradants, matrix components, etc. It can be evaluated by analyzing samples containing the analyte of interest and potential interfering substances.
The tests used to demonstrate specificity will depend on the intended use of the analytical procedure; suitable identification tests should be able to discriminate between compounds of closely related structures, whereas suitable assay or purity tests should be able to discriminate between the analyte, impurities, degradation products and/or matrix components.
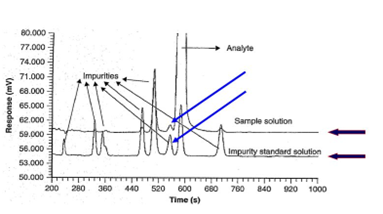
Figure 1: An example of method specificity (ref3)
Linearity
The analytical method should be linear, i.e., there should be a direct relationship between the concentration of the analyte(s) and the signal produced.
Linearity is usually evaluated by analyzing samples containing the analyte at 5 different concentration levels in triplicate. A plot of signals as a function of analyte concentration or content is then created, usually by means of a regression line. The correlation coefficient, y-intercept, slope of the regression line, and residual sum of squares should be calculated.
In case an analytical method does not demonstrate linearity, even after any transformation, the analytical response should be described by an appropriate function of the concentration (amount) of an analyte in a sample.
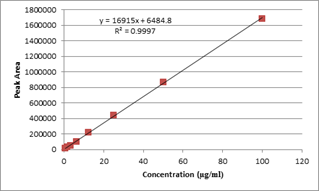 Figure 2: An example of a regression line (ref4)
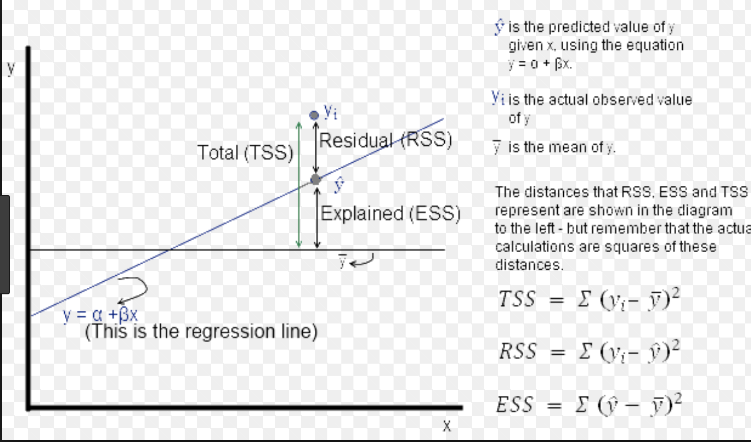
Figure 3: Residuals graph (ref5)
Precision
Precision is the degree of agreement (degree of scatter) between replicate measurements of the same sample under the same conditions.
It is measured by calculating the (relative) standard deviation and confidence interval of the replicate measurements of multiple samplings of the same homogenous sample. Three levels of precision are defined: repeatability, intermediate precision, and reproducibility.
Repeatability: precision established under the same operating conditions over a short of time using a minimum of 6 determinations at 100% of the test concentration or 3 replicates of 3 concentrations.
Intermediate precision: should establish the effects of random events on the precision of the analytical procedure calculated using a minimum of 6 determinations at 100% of the test concentration. Typical within-lab variations such as days, analysts, equipment, etc.
Reproducibility: different analysts in different laboratories should be able to get similar results. This is usually tested in the case of method transfer
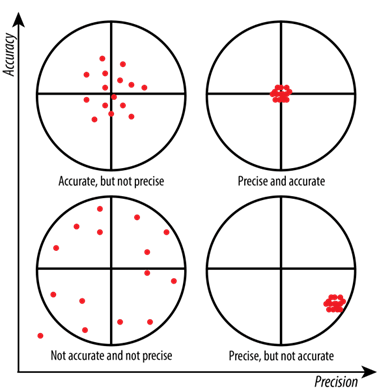
Figure 4: Precision vs Accuracy (ref6)
Accuracy
Accuracy is the closeness of the test results to the true or theoretical value. Accuracy is assessed using a minimum of 9 determinations over a minimum of 3 concentration levels covering the specified range (e.g. 3 replicates each of 3 concentrations over the range). Accuracy is then reported as a percentage recovery of the theoretical amount of analyte in the sample, together with the confidence intervals.
Range
The specified range is normally derived from linearity studies and depends on the intended application of the procedure.
It is established by confirming that the analytical procedure provides an acceptable degree of linearity, accuracy, and precision when applied to samples containing amounts of analyte within or at the extremes of the specified range of the analytical procedure.
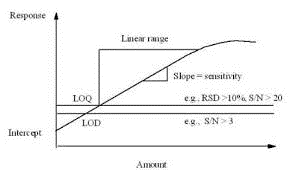
Figure 5: Range (ref7)
The analytical method must be able to detect and quantify low levels of analytes accurately:
- The limit of quantitation (LOQ) is the lowest amount of an analyte that can be quantitated with suitable accuracy and precision.
- The limit of detection (LOD) is the lowest amount of an analyte that can be detected but not necessarily quantitated.
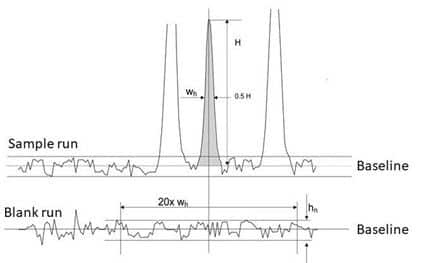
Figure 6: LOQ vs LOD (ref 8)
LOQ and LOD may be calculated using different methods:
- based on visual evaluation,
- based on the Signal-to-Noise ratio,
- or based on the standard deviation of the response and the slope.
The LOQ must be validated by analysis of a suitable number of samples with known concentrations at or near the quantitation limit.
Robustness
The evaluation of robustness should be considered during the development phase. It should show the reliability of an analysis with respect to deliberate variations in method parameters.
An important parameter is the stability of the sample solution.
System suitability testing
System suitability testing is an integral part of many analytical procedures. Tests are based on the concept that the equipment, electronics, analytical operations and samples constitute an integral system that must be evaluated as such. All the (calculated) parameters are within the acceptable limits indicating good performance of the method and system.
Analytical Method Validation guidelines
The go-to guideline for analytical method validation is the ICH Q2. The applicable Pharmacopeias (USP, EP) describe general requirements regarding the use of an analytical technique and might also require certain parameters to be included in System Suitability Testing. These are to be tested prior to each sample run, which includes method validation runs.
Need help validating your analytical methods?
New to Analytical Method Validation? Or do you need advice? Our experts will be happy to help you with the best validation program in accordance with your available resources.
Need help validating your analytical methods?
Our experts will be happy to help you with the best validation program in accordance with your available resources.
Explore our solutions.
| References 1. ICH Topic Q 2 (R1) Validation of Analytical Procedures: Text and Methodology; 1995 2. ICH Q2 Analytical Method Validation (slideshare.net) 3. https://slideplayer.com/slide/6184200/ 4. https://stats.stackexchange.com/questions/265869/confused-with-residual-sum-of-squares-and-total-sum-of-squares 5. Precision Vs. Accuracy – Information Technology (stolaf.edu) 6. http://pharmaceuticalvalidation.blogspot.com/2010/01/validation-of-analytical-methods-and.html 7. HPLC Troubleshooting Tips: Why Signal-to-Noise Ratio Determines the Limit of Detection (analyteguru.com) |
Stay ahead in life sciences
Keeping up with the fast-paced life sciences industry doesn’t have to be overwhelming.
Our newsletter delivers the latest insights, industry updates, and expert content directly to your inbox, helping you stay informed and make smarter decisions.

Discover more expert content

Case study
GMP Support for Closing CAPAs Across Quality Assurance, Qualification & Validation, and Software
Learn how QbD Group helped a biotech CDMO close CAPAs across Quality Assurance, Qualification & Validation, and Software, strengthening GMP compliance and audit readiness.
Whitepaper
Cleaning Validation in Pharma
Learn how risk-based cleaning validation, HBELs, and modern analytical methods help ensure compliant, contamination-free pharmaceutical manufacturing. Download the whitepaper.
Case study
Validating a Complex Autoclave Load to Enhance Sterilization Efficiency
Discover how QbD Group helped a pharmaceutical facility in Barcelona optimize and validate a complex autoclave load, improving sterilization efficiency and manufacturing flexibility.
Case study
Guiding a global biotech company through ATMP facility qualification
Discover how QbD Group supported a global biotech company in commissioning and qualifying two ATMP therapy production sites, ensuring compliance and efficiency.
Case study
Scalable MSAT validation support for seamless compliance & efficiency
Discover how QbD Group helped a global pharmaceutical company address fluctuating validation demands across multiple teams.
Whitepaper
Achieving laboratory compliance
This whitepaper explores the multifaceted aspects of laboratory compliance, including data integrity, quality control measures, and regulatory adherence.
Whitepaper
New GMP Facility Qualification: set-up, process and best practices
This whitepaper delves into the challenges of establishing a new GMP facility, focusing on potential pitfalls and best practices. Download now.
Whitepaper
Analytical Method Validation
In this whitepaper, we will give an overview of the criteria to consider when validating your analytical method.
Case study
Ensuring timely launch: QbD Group's role in establishing a hemophilia drug production line
QbD Group has facilitated the launch of a new drug production line for treating hemophilia, overseeing the qualification of over 100 pieces of small-scale supporting equipment.
Twitter Widget Iframe