
What you need to know to make your medical device UDI-ready in time
What you need to know to make your medical device UDI-ready in time
Regulatory Affairs Medical Devices
Tuesday, 02 March 2021

Maaike Tuinsma

 Share
 \
\
X (Twitter)
 \
\
LinkedIn
 \
\
WhatsApp
 \
\
Copy link
 \
\
Mail
UDI is a new system that assigns a unique barcode to each individual medical device. It will become mandatory when MDR and IVDR enter into force, so as a manufacturer, you need to make sure you’re UDI-ready in time.
Play
What you need to know to make your medical device UDI-ready in time | QbD Group
AI-generated audio
3:47
The EU Medical Device Regulation 2017/745 (MDR) and EU In Vitro Diagnostic Regulation 2017/746, (IVDR)bring new obligations with them. One of them is the UDI system. In this blog post, you will learnwhat theUDI systemis and – more importantly – whyyou should start your own UDI process in time.
What is the consequence of UDI for manufacturers?
The Unique Device Identifier assigns a unique barcode to each individual medical device on the European market. All medical device manufacturers will have to apply the UDI, which will allow importers and distributors to use it for traceability purposes. And regulators, patients, doctors, and others will have easy access to the product information because all UDIs will be recorded in the EUDAMED database.
As a manufacturer, you are responsible for
- the UDI assignment
- the registration in the EUDAMED database
- the integration of the UDI carrier on the label or packaging of the device ( read more about UDI for MDSW here)
The implementation of the UDI system could be challenging. Therefore, we have gathered all relevant information to create a clear overview of the specific UDI requirements.
What exactly is a UDI?
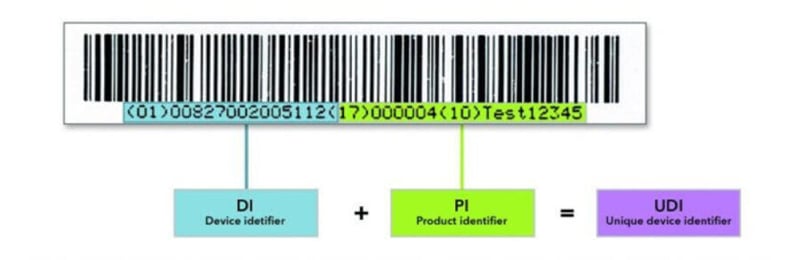
A UDI is a series of numeric or alphanumeric characters specific for a device and manufacturer and comprises the UDI-DIand the UDI-PI.
A UDI-DI(Device Identifier) is required for each individual product and is specific to a version or model of a device, providing access to the information listed in Annex VI Part B. The UDI-DI is therefore static.
The UDI-PI stands for the UDI Production Identifier and identifies the device production unit and, if applicable, the packaged devices, as specified in Annex VI Part C. The UDI-PI is dynamic as it varies with the production characteristic of the product. It gives information about the lot number, serial number, manufacturing date, expiration date, etc.
To obtain a UDI-DI and UDI-PI code, you need to contact one of the entities the European Commission authorizes. The information that needs to be provided depends on the chosen entity.
The Basic UDI-DI and the UDI carrier
The Basic UDI-DI is an identification number specific to a group of devices with the same intended purpose, risk class, and essential design and manufacturing characteristics of a manufacturer. It is independent of the packaging or labeling of the device and does not appear on any trade item.
The Basic UDI-DI is the main access key for device-related information in the EUDAMED database and it is provided by officially designated entities selected by the European Commission. How the Basic UDI-DI is generated, the format it comes in, and the information that needs to be provided depends on the entity you choose to provide the Basic UDI-DI.
The UDI carrier is an expression of the representation of the UDI. It consists of an Automatic Identification and Data Capture (AIDC) part and a Human Readable Interpretation (HRI) part. The AIDC is a technology used to automatically capture data, including bar codes, smart cards, biometrics, and radio-frequency identification (RFID). The HRI is a legible interpretation of the data characters encoded in the UDI. Both the AIDC and HRI should contain the complete UDI information.
The UDI carrier must be placed on the label of the device and on all higher levels of packaging as well. In the case of reusable devices, it must be placed on the device itself (direct marketing). Some exceptions and more specific requirements concerning the UDI carrier can be found in Annex VI Part C Section 4 of the MDR/IVDR.
When do you have to be UDI-ready?
The UDI requirements will become mandatory once the MDR (26 May 2021) and IVDR (26 May 2022) enter into force. UDI data submission in the EUDAMED database will become mandatory on 26 November 2022, which will be 24 months after EUDAMED became fully functional. Concerning the UDI carrier, it depends on the classification of your device:
- 26 May 2021 for Class III and implantable devices
- 26 May 2023 for Class IIa and IIb and Class D devices
- 26 May 2025 for Class I and Class B and C devices
- 26 May 2027 for Class A devices
What if I’m not UDI-ready in time?
So as you can see, depending on the type of device you produce, you may not have much time left to do your UDI homework. That’s why we strongly advise you to start your UDI process in time.
In case additional support is needed, we kindly invite you to talk to our QbD experts.
Need regulatory support?
Trust our expertise to guide you through the regulatory process and achieve compliance
with ease.
Discover how we can help.
Stay ahead in life sciences
Keeping up with the fast-paced life sciences industry doesn’t have to be overwhelming.
Our newsletter delivers the latest insights, industry updates, and expert content directly to your inbox, helping you stay informed and make smarter decisions.

Discover more expert content
Webinar
From Strategy to First-in-Human: A Practical Roadmap for Biotech Development
Join our webinar to learn how to navigate nitrosamine risks and streamline your biotech development from early planning to first-in-human trials. Register now for expert insights.
Webinar
PMS & PMCF Compliance for Medical Devices Under EU MDR
Gain practical insights on PMS & PMCF compliance for medical devices under EU MDR. Watch our webinars on demand.
Case study
A Decade of Excellence: Linguistic Review Support for a Global Pharma Leader
QbD Group streamlines linguistic reviews for a leading pharma client, ensuring compliance and accuracy across 25 EU/EEA languages while meeting stringent regulatory deadlines.
Case study
Supporting Market Access to Essential Medicines in Europe
Support your entry into the European pharmaceutical market with expert regulatory guidance, ensuring compliance and access to essential medicines for patients. Contact us today for assistance.

Whitepaper
Medical Device Regulation (MDR) Checklist
Implement the Medical Device Regulation (MDR) with ease. Download our checklist of mandatory documents for MDR compliance.
Whitepaper
Regulatory Affairs for Pharma and Biotech
In this flyer, you will learn more about the regulatory services QbD Group provides for the pharmaceutical and biotechnology industries.
Whitepaper
Clinical Evaluation for Medical Devices under MDR
This whitepaper will guide you through crucial regulatory documents pertaining to the clinical evaluation process of your medical device. Download now!
Whitepaper
IVDR extension: what does this mean for you?
In this whitepaper, you will learn the impact of the IVDR transition extension and receive tips and strategies to navigate these regulatory changes.
Webinar
Drug-Device Combination Products and Article 117
Explore the regulatory complexities of Drug-Device Combination Products. Learn about Article 117 requirements, Notified Body Opinions, and more.
Whitepaper
Mobile health on the rise: exploring the regulatory landscape for reimbursement
This whitepaper will help you navigate the maze of the DTx regulatory environment, highlighting several important countries and regulations.
Whitepaper
Clinical investigations under MDR: key insights and practical experiences
This whitepaper covers clinical investigation authorization under MDR, the EU Legal Representative's role, and the MDR's impact on study conduct and post-market surveillance.
Case study
CE Mark Renewal for bioXtra Products: A Success Story
Discover how Lifestream Pharma successfully renewed the CE mark for their BioXtra Dry Mouth products under the EU MDR. Learn about the challenges faced, the approach taken for compliance, and the results achieved.
Webinar
Technical Documentation Essentials for Medical Device Software
Gain practical insights on crafting MDR-compliant technical documentation for Medical Device Software.

Webinar
Getting Started: Overcoming Initial Obstacles in Medical Device Software Development
Watch our webinar on demand and learn about regulatory obstacles, MDR, AI Act, and best practices for medical device software development and market entry.
Webinar
Mastering Clinical Performance Studies under IVDR
In the evolving regulatory landscape of IVDs, demonstrating clinical performance is more crucial and challenging than ever. Discover more in this webinar on demand.
Whitepaper
Clinical evidence for In Vitro Diagnostics under IVDR
In this whitepaper, we will guide you through the documents required for the demonstration of clinical evidence under the IVDR. Download now.
Webinar
Post-Market Surveillance Requirements for Medical Devices and IVDs
Learn about post-market surveillance requirements under MDR and IVDR regulations. Gain insights from industry experts in this on-demand webinar.
Whitepaper
The key role of Regulatory Affairs in the pharmaceutical industry: from drug development to commercialization
Discover the key role of Regulatory Affairs in pharma and how Regulatory Affairs teams support product lifecycle management in this whitepaper. Download now.

Webinar
3D Printing in the Healthcare World
Explore the challenges and opportunities of 3D printing in healthcare, featuring expert insights on regulations, pitfalls, and efficiency.

Webinar
IVDR Extension Explained: Tips & Strategies
Gain valuable insights on the IVDR extension and its impact on IVD manufacturers with our webinar on demand.
Whitepaper
Innovations in ATMP: state of the industry in 2024
This whitepaper explores the innovations in ATMP and solutions in detail, with a focus on the current landscape as of April 2024. Download now.

Case study
The journey toward IVDR compliance for the LVOne device
QbD Group helped Upfront Diagnostics gain entry into the MD/IVD industry by supporting them on their path to ISO 13485 certification.
Case study
Ensuring a smooth MDR transition for Oystershell's medical devices
The landscape of medical device regulations is constantly evolving, making it a challenge for companies to keep up. Our collaboration with Oystershell, which began in July 2023, highlights our expertise in transforming complex challenges into success stories, all with a touch of QbD Clinical magic.
Case study
Successful clinical evaluation supports rapid CE marking of Minze Health's Uroflowmeter
QbD Clinical assisted Minze Health, a urology-focused medical devices company, in obtaining CE marking under MDR through clinical evaluation.

Webinar
The Medical Device CE Certification Pathway Explained: From Classification to Conformity Assessment by the Notified Body
Learn about the Medical Device CE Certification Pathway, from classification to conformity assessment by a Notified Body. Join our webinar on demand.

Case study
Helping Relu to comply with changing regulations
QbD Group supported software company Relu with their expertise to comply with changing regulations.

Case study
Quickly bringing a new COVID-19 medical device to market
Obtaining the required government certifications for a medical device is always a challenge. Especially when the COVID clock is ticking.
Twitter Widget Iframe